
Retain Responsibility Where Activities Are Transferred
The sponsor board should ask management how it establishes the reliability of material evidence produced by a contract research organization or other AI-enabled supplier. Delegating an activity can change who performs the work without changing who must answer for the trial or regulatory submission.
ICH E6(R3) states that a sponsor can transfer trial activities while retaining ultimate responsibility for those activities and trial-data reliability.1 Regional adoption and applicable requirements must be checked. The guideline is not a universal direct approval of any AI method or supplier.
Identify the trial activity, intended AI role, source data, responsible qualified people, and evidence entering the sponsor’s decision. Distinguish drafting, extraction, analysis, interpretation, and submission. Each can create a different dependency and require different verification.
Govern the Evidence Pipeline Across the Supplier Chain
The provider may use further model, data, technology, or human service providers. The sponsor needs sufficient access and oversight to understand material contributions. An agreement with the prime supplier should lead to a usable evidence route through the actual delivery chain.
The study reports downstream dependency gaps and manual alterations in banking delivery.2 It does not establish clinical-trial data defects. The transferable concern is whether the sponsor can see the work and changes behind a final artifact it is expected to rely on.
For a hypothetical trial summary, a fluent final document may contain extracted results, model-generated wording, and human corrections. Preserve the sources, material transformations, review decisions, and final approved version. A statement that a human reviewed the output should identify what that review established.
Use a Sponsor Evidence Responsibility Record
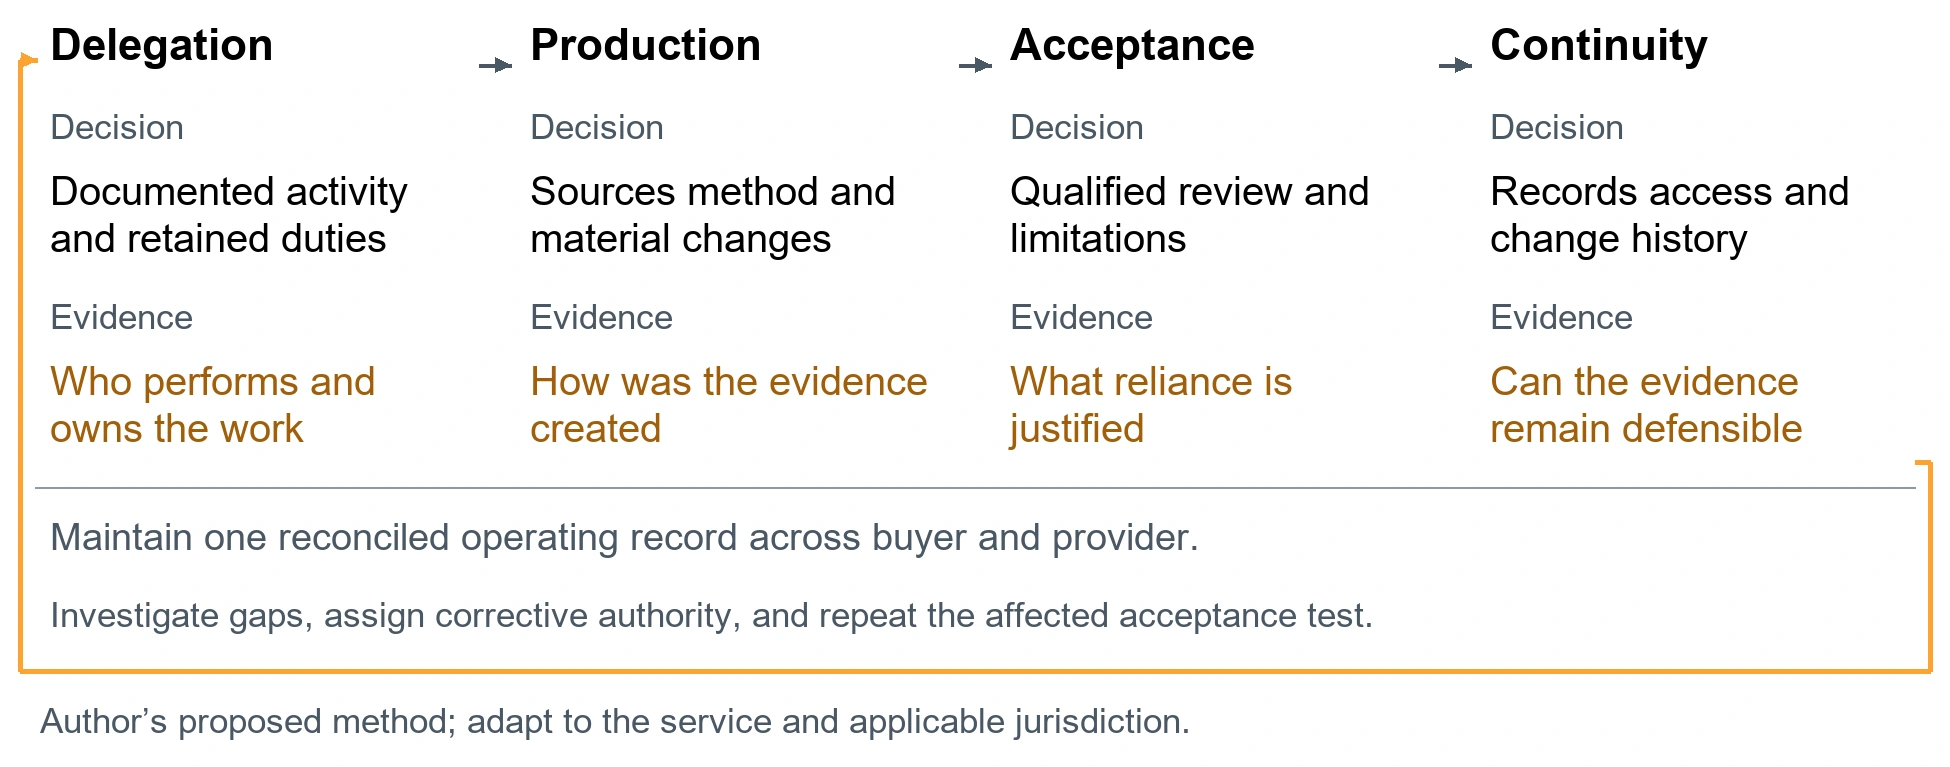
Figure 1 proposes four links: delegated activity, evidence production, qualified acceptance, and continuing access. Record the sponsor owner, provider owner, approved method, relevant configuration, source provenance, and conditions that require renewed review.
FDA and EMA’s January 2026 principles emphasize clear context of use, traceable data governance, and lifecycle assessment.3 They are guiding principles, not product authorization. The sponsor should apply the relevant requirements and evidence standards to the specific trial activity.
Tests should assess the complete workflow, including human-AI interaction and corrections. Define when the sponsor can rely on provider quality work and what independent challenge remains necessary. Where an important claim cannot be substantiated, restrict its use rather than convert uncertainty into a favorable acceptance statement.
Preserve Inspection and Change Capability
EU, UK, Swiss, US, and other clinical-trial and medicines requirements differ in implementation and activity. Management should establish applicable GCP, privacy, records, validation, and submission expectations. Do not assume one model evaluation satisfies all trial or regulatory evidence needs.
Contracts should support access to relevant records, personnel, audit trails, and downstream information for the required period. Plan for provider exit or platform retirement. The sponsor’s duty to explain historical evidence can outlast the commercial service that produced it.
A change to a model, extraction method, data source, or human review process can affect the meaning of the evidence. Require notice and an assessment of the affected validation and acceptance basis. Preserve the configuration and record applicable to each material output.
Require Demonstrable Sponsor Oversight
A reasonable objection is that sponsors cannot duplicate all supplier work. Oversight should be risk based and proportionate. The sponsor can rely on suitable provider controls where justified while retaining enough evidence and expertise to challenge consequential claims and investigate exceptions.
Commission a targeted reconstruction of important outsourced evidence before material reliance. Trace it from source through transformation, intervention, review, and sponsor acceptance. Include a changed or failed case. Resolve gaps with qualified clinical, quality, statistical, legal, and technical owners as appropriate.
The board should approve an outsourcing model that preserves the sponsor’s ability to substantiate the evidence it uses. A capable supplier can strengthen development. The sponsor’s responsibility remains credible only when the evidence, oversight, and continuing access survive the two-party delivery model.
Evidence and Source Note
The study is a nonrepresentative cross-section of 850 vendor delivery-team respondents on 150 engagements serving Tier 1 financial institutions, collected during January–June 2026. Neither banks nor vendors corroborated the accounts. It establishes reported experience, not causation, legal breach, or market prevalence. Its application here is a mechanism to investigate, not an estimate of conditions in other industries. The author-supplied version 1.8.2 is dated November 1, 2026 and is cited as a forthcoming manuscript consulted October 23, 2026.4
This is an analytical decision paper based on the supplied study and selected primary public sources checked through October 23, 2026. The original contribution is a sponsor evidence-responsibility record connecting delegated AI work to provenance, qualified reliance, and durable inspection access. The proposed method has not been independently validated. Legal requirements depend on the entity, activity, jurisdiction, and effective provisions; supervisory observations, research, and professional commentary have distinct authority.
Notes
- International Council for Harmonisation of Technical Requirements for Pharmaceuticals for Human Use, Guideline for Good Clinical Practice E6(R3) (International Council for Harmonisation of Technical Requirements for Pharmaceuticals for Human Use, January 6, 2025), Annex 1, secs. 2.12.2, 3.6.4–3.6.10, and glossary definition of audit trail, https://database.ich.org/sites/default/files/ICH_E6(R3)_Step4_FinalGuideline_2025_0106.pdf. ↩
- Phil Hatch, Agentic AI Outsourcing at Tier 1 Financial Institutions: Full Study Report, version 1.8.2 (Akholi, forthcoming manuscript dated November 1, 2026), Part 3, Third-Party Ecosystem, sec. 6.1, “Mapping of Third and Fourth Party Dependencies,” indicator C196, https://doi.org/10.2139/ssrn.7164559; Hatch, Agentic AI Outsourcing, Part 5, “The Five Manual Alteration Methods” and “Disclosure of Manual Alteration,” table 5.8. ↩
- US Food and Drug Administration and European Medicines Agency, Guiding Principles of Good AI Practice in Drug Development (US Food and Drug Administration and European Medicines Agency, January 2026), principles 2, 4–6, 8–10, https://www.fda.gov/media/189581/download. ↩
- Hatch, Agentic AI Outsourcing, Introduction, “Executive Summary” and “Evidentiary Basis and Its Limits”. ↩
Bibliography
- Hatch, Phil. Agentic AI Outsourcing at Tier 1 Financial Institutions: Full Study Report. Version 1.8.2. Akholi. Forthcoming manuscript dated November 1, 2026. Author-supplied manuscript consulted October 23, 2026. https://doi.org/10.2139/ssrn.7164559.
- International Council for Harmonisation of Technical Requirements for Pharmaceuticals for Human Use. Guideline for Good Clinical Practice E6(R3). International Council for Harmonisation of Technical Requirements for Pharmaceuticals for Human Use, January 6, 2025. https://database.ich.org/sites/default/files/ICH_E6(R3)_Step4_FinalGuideline_2025_0106.pdf.
- US Food and Drug Administration and European Medicines Agency. Guiding Principles of Good AI Practice in Drug Development. US Food and Drug Administration and European Medicines Agency, January 2026. https://www.fda.gov/media/189581/download.
Explore this subject
Category: Regulation and Compliance
Topics: Evidence and Traceability Retained Accountability Safety and Validation
Audience: Buy-side Boards and Executives
Industry: Life Sciences Other Regulated Industries